Sterilisation Methods: Steam and Dry Heat (Ph. Eur. 5.1.1)
Contents
- The general idea
- Steam (moist heat) sterilisation
- Principle
- Equipment
- Sterilisation cycle
- Cycle effectiveness
- Routine control
- Dry heat sterilisation
- Principle
- Equipment
- Sterilisation cycle
- Cycle effectiveness
- Routine control
- Other methods in brief
- Frequently asked questions
- What is the difference between sterilisation and depyrogenation?
- Why are biological indicators still needed if temperature is recorded?
- Can we use a different time and temperature from the reference cycle?
- How long must sterilisation records be kept?
- See also
- References
In short: European Pharmacopoeia general chapter 5.1.1 describes the accepted methods of sterilisation. This article explains, in our own words, how steam (moist heat) and dry heat sterilisation work, what the equipment and cycle must achieve, and how the process is validated and monitored. It also shows where EU GMP Annex 1 (2022) covers the same topics. For the exact wording and limits, read the current Ph. Eur. text.
The general idea
A sterilisation method may be modified or combined with another, provided the chosen process is validated for two things: its effectiveness in killing micro-organisms, and the integrity of the product and its container. During every cycle the critical parameters are monitored to confirm that the conditions defined in validation are met throughout the batch. This applies even when the standard reference conditions are used.
Related Ph. Eur. chapters: 5.1.2 (biological indicators of sterilisation) and 5.1.5 (the F0 concept for steam sterilisation). After sterilisation, protect the items from recontamination. For the overall expectations on sterile products, see EU GMP Annex 1 (2022), Section 8, “Production and specific technologies”: terminal sterilisation is preferred where the product allows it (paragraph 8.34), and the sterilisation process must be chosen, designed and validated on scientific data (paragraph 8.35).
Steam (moist heat) sterilisation
Annex 1 (2022), paragraph 8.55, describes moist heat as steam in direct or indirect contact with the load, and also superheated water systems for containers that other cycles could damage.
Principle
Steam gives up its heat when it condenses on cooler surfaces. For items in direct contact with steam, the steam must reach every surface, which means air and other non-condensable gases must be removed. The moisture also helps kill organisms. For sealed containers, the autoclave chamber only acts as a heating jacket: the kill happens from the temperature reached inside the container, in the product and in the head-space.
Equipment
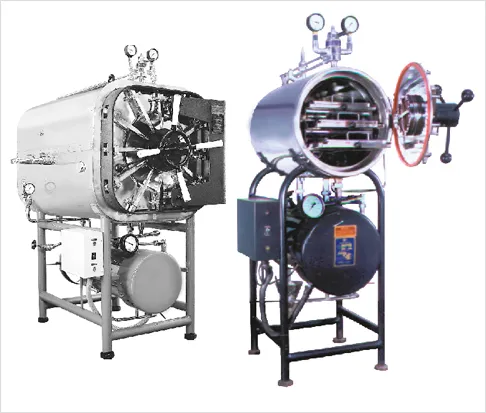
Steam sterilisation is carried out in an autoclave, a pressure vessel that admits or generates steam and removes condensate to hold the required pressure and temperature. Key points:
- The steam supply for direct-steam cycles should be saturated and free of non-condensable gases.
- For sealed containers, steam-air mixtures or a superheated water spray can be used to transfer heat and to balance pressure inside containers.
- The autoclave must be qualified for uniform conditions in the chamber and in the load, and for the loading patterns used. Temperature profiles are recorded in the slowest-to-heat items.
- Chamber temperature and pressure sensors must be suitable and placed in relevant positions, and profiles are recorded for every cycle. At least one independent probe should follow the load at the slowest-to-heat position.
- Water used to cool sealed containers at the end of the cycle must be of a quality that does not harm sterility.
Sterilisation cycle
The cycle has to suit the items and the loading pattern. Where air leaves by gravity displacement, items must be arranged so no air pocket is trapped. Where air is removed by vacuum and steam pulses, the items must tolerate evacuation. Porous loads and hollow items need cycles that remove air reliably. Steam penetration is checked during cycle development, for example with physical or chemical indicators, and biological effectiveness is confirmed with biological indicators. Loading patterns must be specified.
Cycle effectiveness
The Ph. Eur. reference condition for steam sterilisation is 15 minutes at 121 °C in saturated steam, measured in the coldest position of the chamber. Other time-temperature combinations are allowed if developed and validated for the product and load. The EMA sterilisation guideline (2019) adds that every steam sterilisation process needs a minimum lethality of F0 8 minutes and a minimum hold temperature of 110 °C.
F0 is the lethality delivered by a cycle expressed as equivalent minutes at 121 °C (see Ph. Eur. 5.1.5). The physically calculated value (Fphys) is correlated with the biological result (Fbio), which is derived from the D-value at 121 °C of the biological indicator and the drop in its viable count:
Fbio = D121 × (log N0 − log N)
where N0 is the count before exposure and N the count after exposure.
During validation, find the positions in the load that are hardest to sterilise and place biological indicators there (or in product). Allow for protection of spores, for example by steam occlusion or by the product itself. The Fbio at the hardest position defines the parameters needed to achieve the required sterility assurance level (SAL) of 10-6 or better. Annex 1 (2022) expects probe positions for control and recording to be justified in validation (paragraph 8.51), validation of porous cycles to cover equilibration time, exposure time and pressure-temperature correlation (paragraph 8.59), and, for fluid loads in superheated water autoclaves, temperature mapping of the entire load, heat-penetration and reproducibility studies, with routine probes correlated with the worst-case positions (paragraph 8.65).
Routine control
For each cycle, record chamber pressure, temperature and time, at least in the coldest position of the chamber, and calculate and record F0 where possible.
Dry heat sterilisation
Principle
Dry heat is a terminal method in which heat is passed to the articles by convection, radiation or direct contact. Annex 1 (2022), paragraph 8.66, notes it is particularly useful for removing thermally robust contaminants such as endotoxin (pyrogen), and is often used to prepare components for aseptic filling.
Equipment
A forced-air oven or other purpose-built equipment such as a tunnel. For tunnels, airflow must protect the grade A sterilising zone, with suitable pressure differences (paragraph 8.67). Ovens should stay at positive pressure relative to lower-grade areas during sterilisation and the hold afterwards (paragraph 8.70).
Sterilisation cycle
Load the steriliser so that the required temperature is reached throughout the load. Place temperature sensors in or on representative items in the coolest position, which is established beforehand. Record time and temperature for every cycle.
Cycle effectiveness
The Ph. Eur. reference condition is at least 160 °C for at least 2 hours, but other time and temperature combinations may be used when shown to give adequate, reproducible lethality within the established tolerances.
Validation combines temperature mapping with biological indicator studies. For depyrogenation of glassware, dry heat above 220 °C for a validated time is common; a 3 log10 reduction of heat-resistant endotoxin can be used as the validation criterion (EMA sterilisation guideline, section 4.1.2). Annex 1 (2022), paragraph 8.69, requires endotoxin-spiked containers during validation.
Routine control
Record temperature profiles at least in the coldest position of the chamber, and time and temperature for each cycle.
Other methods in brief
Annex 1 (2022) also covers the following, which the Ph. Eur. describes in later parts of chapter 5.1.1:
- Ionising radiation is used mainly for heat-sensitive materials and products. Ultraviolet irradiation is not an acceptable sterilisation method (paragraph 8.71, with guidance in Annex 12).
- Ethylene oxide (gas) should be used only when no other method is practicable, with proof of no damage to the product and of adequate degassing to residue limits (paragraph 8.73).
- Sterilising filtration uses a sterilising-grade filter, with integrity testing before use (PUPSIT, where possible) and after use (paragraph 8.87) and, for liquids, normally one filter per batch (paragraph 8.94).
Frequently asked questions
What is the difference between sterilisation and depyrogenation?
Sterilisation destroys viable micro-organisms. Depyrogenation inactivates or removes endotoxin, which survives normal sterilising conditions. Dry heat at high temperature is the usual way to depyrogenate glass.
Why are biological indicators still needed if temperature is recorded?
Temperature records show what the equipment delivered. Biological indicators show that the delivered conditions actually kill a resistant challenge organism at the hardest-to-sterilise position.
Can we use a different time and temperature from the reference cycle?
Yes, if the alternative is developed and validated for the specific product and load and delivers the required SAL, as the chapter allows.
How long must sterilisation records be kept?
Keep cycle records as batch-related GMP records for at least 1 year after the expiry date of the respective product batch (EU GMP Part I, Chapter 4, paragraph 4.11; 21 CFR 211.180(a)), or longer if local law or the marketing authorisation requires.
See also
References
- European Directorate for the Quality of Medicines (EDQM), European Pharmacopoeia, general chapter 5.1.1 Methods of preparation of sterile products, and chapters 5.1.2 and 5.1.5 (current edition; text not freely accessible).
- European Commission, EudraLex Volume 4, Annex 1: Manufacture of Sterile Medicinal Products (revision published 2022), Section 8, paragraphs 8.34, 8.35, 8.50 to 8.73, 8.87 and 8.94.
- European Medicines Agency, Guideline on the sterilisation of the medicinal product, active substance, excipient and primary container, EMA/CHMP/CVMP/QWP/850374/2015 (March 2019), sections 4.1.1 and 4.1.2 (quotes the Ph. Eur. 5.1.1 reference conditions).
- European Commission, EudraLex Volume 4, EU GMP Part I, Chapter 4: Documentation, paragraph 4.11; 21 CFR 211.180(a).
Related articles
SOP for Microbial Culture Management in a QC Laboratory
Example SOP for microbial culture management: procurement, reconstitution, seed lot and working cultures, passage limits, purity checks, storage and disposal.
Microbiologist Qualification SOP (Example Template)
Example SOP for qualifying QC microbiologists: training, blind-coded samples for MLT, sterility, BET and LBPC, reviewer checks, records and requalification triggers.
SOP: Entry, Exit and Gowning in Non-Sterile Microbiology Lab
Example SOP for entry, exit and gowning in a non-sterile microbiology testing area: airlock steps, health precautions, gown reuse rules, records and training.
SOP on Laminar Air Flow (LAF): Operation, Cleaning, Qualification
Example SOP for a laminar air flow (LAF) workstation in a microbiology lab: operation, cleaning, pre-filter care, air velocity, particle count and HEPA integrity tests.